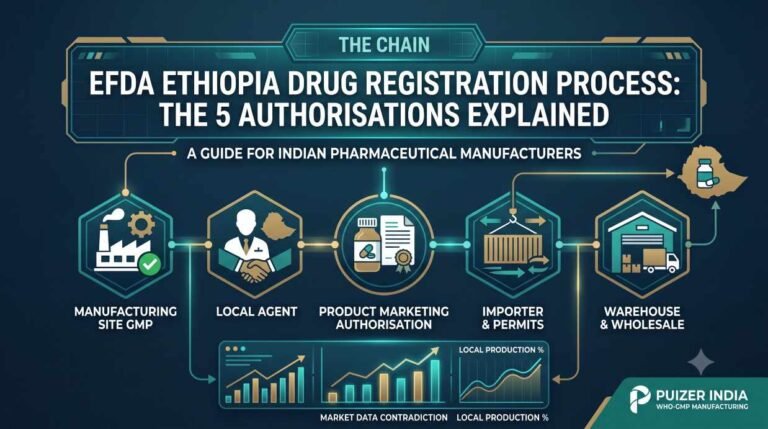

What the EFDA drug registration process requires
The EFDA drug registration process ends in a marketing authorisation from the Ethiopian Food and Drug Authority, issued under Article 20 of the Food and Medicine Administration Proclamation No. 1112/2019[1] and applied for online through i-Register on EFDA’s eRIS portal.[2] A foreign manufacturer files through a local agent physically located in Ethiopia, whose pharmacist technical person holds the eRIS account.[3]
The dossier follows the CTD format in English, as editable PDF. For a multisource generic, Modules 4 and 5 are replaced by the bioequivalence study report.[3] EFDA notifies acceptance or deficiencies within 15 days, gives six months to answer queries, and issues the certificate only after the manufacturing site has cleared GMP inspection or an EFDA GMP waiver.[3] The authorisation is valid for five years; renewal is filed within the 180 days before expiry.[4]
Scope of this guide: getting a product registered. Importing it after registration, whether to EPSS tenders or a private importer, with import permits and port clearance, is covered in how to import medicines from India to Ethiopia.
EFDA drug registration process in 8 steps
Ethiopia is benchmarked by WHO at Maturity Level 3 for medicines and imported vaccines since 30 September 2025, which WHO describes as a stable, well-functioning and integrated regulatory system.[5] Ethiopia was the ninth African country at ML3 when WHO announced it; Mozambique became the tenth in August 2026.[6] In practice ML3 means the steps below are applied as written, so plan for them rather than around them.
-
Manufacturer
Confirm the registration route
The route decides the timeline. A WHO-prequalified product can use the collaborative procedure; a product approved by one of the reference authorities on EFDA’s reliance list can use an abridged review; a product approved only in India takes the full assessment.[7] Use the route selector further down this page.
-
Manufacturer + local agent
Appoint the local agent and sign the agency agreement
The agent must be physically located in Ethiopia. The agreement is signed and stamped by both parties and uploaded with the application.[3] The agent is your registrant, so choose it before you compile anything.
-
Local agent
Register the technical person on eRIS
EFDA issues eRIS credentials only to the technical person the agent or manufacturer assigns. That person must be a pharmacist and must be registered with EFDA.[3]
-
Manufacturer
Compile the CTD dossier
English, editable PDF (scans only for certificates), A4, Times New Roman 12 point.[3] The module-by-module contents are in the tabs below.
-
Local agent to EFDA
Submit on i-Register and pay the service fee
Submission is online only. eRIS issues a verification number, and applications are assessed first-in, first-out by that number, except fast-track and non-routine applications.[3] The fee is set by the Rate of Service Fees Regulation No. 370/2015.[8]
-
EFDA
Screening within 15 days
EFDA notifies acceptance or deficiencies within 15 days of submission, unless workload or a non-routine procedure prevents it.[3]
-
EFDA + applicant
Assessment and queries
Queries must be answered within six months of notification. After that EFDA sends a 15-day request to supplement, then rejects the application if nothing arrives.[3]
-
EFDA + manufacturer
GMP clearance, then the certificate
No inspection is needed before you file, but the site must be inspected, or covered by an EFDA GMP certificate or waiver letter, before the marketing authorisation is issued.[3] How EFDA decides between on-site, desk and reliance clearance is explained in the GMP section below.
EFDA local agent requirements for foreign manufacturers
EFDA’s registration guideline requires every foreign manufacturer that imports or offers medicines for import into Ethiopia to appoint a local agent or representative in Ethiopia.[3] Four clauses decide how much control you keep over your own registration.
A pharmacist runs the file
The agent’s technical person should be a pharmacist, and only that person receives the eRIS login.[3] If they leave, notify EFDA and reassign the account, or a live application goes silent.
One registrant, more distributors only if written in
More than one distributor is allowed only when the agency agreement says so.[3] Draft regional distribution rights at the start; adding them later means a new agreement.
Shared recall and pharmacovigilance
The parties to the agreement are responsible for recall if fraud or an unacceptable adverse event occurs, and both share pharmacovigilance and post-marketing reporting.[3]
No exit before approval without cause
The agreement cannot be terminated without a justifiable reason before the marketing authorisation is granted.[3] Check the agent’s track record before you sign, not after.
The same guideline says the agent cannot import the registered product without a valid EFDA certificate of competence and a trading licence.[3] Whether you need an agent that also imports depends on who buys your product. That choice, EPSS tender or private importer, is mapped in our India to Ethiopia import guide.
Which EFDA registration route does your product qualify for?
EFDA runs a full assessment, two reliance reviews, the WHO collaborative procedure, a fast-track queue and a non-routine track. Answer three questions.
Indicative routing from published EFDA guidelines. EFDA confirms eligibility at screening.
The routes side by side
| Route | Who it is for | What EFDA reviews | Published timeline | Instrument |
|---|---|---|---|---|
| Full (routine) assessment | No qualifying reference approval, e.g. India-only generics | CTD dossier; BE study report for multisource generics[3] | Acceptance or deficiency notice within 15 days; no end-to-end day count published[3] | EFDA/GDL/017 v005 |
| Fast-track | HIV/AIDS, malaria, TB, vaccines, maternal and child health, anti-cancer, hepatitis, new drugs, orphan drugs, locally produced medicines[3] | Same technical requirements as routine | Priority in the queue, not a lighter review | EFDA/GDL/017 v005 |
| Verification review (reliance) | WHO-prequalified products; products accepted through AMA or IGAD joint assessment[7] | Administrative check that the product is the same as the one assessed | No day count in the guideline; refers to Directive No. 963/2023[7] | EFDA/GDL/042 v003 |
| Abridged review (reliance) | Products approved by an authority on EFDA’s Annex I list of 39 entries; India’s CDSCO is not on it[7] | Only the dossier sections relevant to Ethiopia | No day count in the guideline[7] | EFDA/GDL/042 v003 |
| WHO collaborative registration | WHO-prequalified products, with the holder’s consent to information sharing[9] | Dossier as accepted by WHO PQ, with WHO assessment and inspection outcomes | Certificate within 90 days once documents are fulfilled[9] | EFDA/GDL/021 v002 |
| Non-routine: conditional approval or emergency use | Emergencies, verified unmet need, listed orphan medicines, promising Phase III products with no alternative[10] | Task-force assessment; data requirements may be reduced | Report within a week; steering committee decision within 3 days; conditional approval valid up to 1 year[10] | EFDA/GDL/024; Directive 963/2023 Arts 23 and 24 |
On a phone, scroll the table sideways.
EFDA publishes a 90-day commitment for the WHO collaborative route and working-day windows for renewal, but no end-to-end day count for a full assessment. Any quote of “EFDA takes X months for a normal generic” is an anecdote. Plan against the control points you can check: the 15-day screening notice, the six-month query window and GMP clearance before the certificate.[3]
EFDA CTD dossier requirements, module by module
EFDA runs three e-services: i-Register for product registration, i-Import for import permits and i-License for certificates of competency.[2] The dossier goes into i-Register. The online form and the uploaded files must match; the applicant is responsible for that consistency.[3]
Module 1: administrative and product information
The Ethiopia-specific module, and the one that fails most often because every document in it is dated:
- eRIS application form and a dated covering letter
- Agency agreement, signed and stamped by both parties[3]
- Valid GMP certificate or manufacturing licence from the exporting country’s authority
- Certificate of Pharmaceutical Product (CoPP), or instead a valid cGMP certificate plus the marketing authorisation from the exporting country’s regulator; WHO prequalification and certificates from WHO-listed authorities are considered case by case[3]
- SmPC, labels and patient information leaflet
- Proof of the service fee payment[3]
What each certificate is and who issues it in India is set out in our pharmaceutical export documentation checklist.
Module 2: summaries
The common CTD overviews and summaries: quality overall summary and, where Modules 4 and 5 apply, the non-clinical and clinical overviews. Modules 2 to 5 follow the common CTD structure; only Module 1 is authority-specific.[3]
Write the quality summary from the same data you file in Module 3. A summary that cites a different batch, site or specification than Module 3 is a standard query trigger.
Module 3: quality
API and finished-product data: manufacture, controls, specifications, validation, container closure and stability. For an Indian exporter this is where most of the work sits. Stability data must support the pack and climate the product will actually meet; see climate-zone stable packaging for export.
If you plan a reliance route, keep Module 3 identical to the reference dossier. EFDA’s sameness test covers formulation, API and finished-product sites (down to the block or unit), processes, and specifications.[7]
Modules 4 and 5: non-clinical and clinical
For a multisource generic where a bioequivalence study is mandatory, Modules 4 and 5 are not applicable, except that the BE study report is filed with the other modules.[3] New products file full non-clinical and clinical modules.
Check the BE requirement for each product before you price the registration; the BE study, not the dossier compilation, is usually the largest cost and the longest lead time.
Labels and leaflets
- Label statements in English and/or Amharic.[3]
- For medicines on the Essential Medicines List, or in wide circulation, the patient leaflet must be at least in English and Amharic.[3] Check your products against the current list.[11]
- EFDA issued a Medicine Packaging and Labeling Directive, No. 1091/2025, in June 2025.[12] Its text is published mainly in Amharic and we have not summarised it here; ask your agent to check artwork against it before you file.
EFDA GMP inspection of an Indian manufacturing site
GMP clearance is the step that most often holds a finished assessment. EFDA’s Medicine GMP Inspection Procedure Directive No. 1055/2025 sets how a foreign site is cleared.[13] (The Amharic header numbers it 1055/2017: that is the Ethiopian-calendar year.)
Who can be inspected
A foreign site is eligible only if its home authority licenses it to make medicines and it has produced continuously for at least one year.[13]
On-site, desk or reliance
A routine on-site inspection is the default for new facilities. Remote or desk review can replace it after a risk assessment, and EFDA may waive on-site inspection for companies approved by a reference authority, with random confirmatory visits.[13]
When it happens
On-site inspection normally follows completion of dossier evaluation, though it can come earlier. Payment must be effective first, and the site gets at least two weeks’ notice.[13]
How long the result takes
With no deficiencies, the certificate follows within 10 working days of the last inspection day; deficiencies trigger a CAPA cycle with 20- and 10-day steps. Inspection frequency is at most three years.[13]
India’s CDSCO is not one of EFDA’s reference authorities,[7] so on the directive’s wording the reliance waiver does not apply to a site whose only approvals are Indian. Plan for an on-site inspection, or a desk review if EFDA’s risk assessment allows it, and budget its fee: our registration fee comparator sets EFDA’s inspection fee beside seven other African regulators. Correspondence must be in English or carry a certified translation.[13] The batch evidence inspectors and importers ask to see is covered in what a WHO-GMP medicine supplier should show.
EFDA marketing authorisation renewal
A marketing authorisation runs five years. Renewal follows EFDA’s Guideline for Renewal of Marketing Authorization, EFDA/GDL/018 version 003:[4]
- When: file within the 180 days before expiry, through eRIS.
- If late: a further 180-day grace period at the new-application fee. Miss that and the authorisation is treated as revoked; you file a whole new dossier with technical data.
- What: administrative documents, valid GMP certificate and licence, the agency agreement or a no-change declaration, periodic safety information, a table of post-marketing variations and the fee receipt. Technical modules are not re-filed.
- How long: about 60 working days, not more than 90 working days from submission, with clock stops for queries.
Where EFDA registrations lose time
- Choosing the agent last. The eRIS account and the agency agreement both sit before your first upload.[3]
- Treating GMP clearance as a formality. A dossier that clears assessment and then waits for an inspection slot has wasted its place in the queue.[13]
- Burning the six-month query window. An API source change or a stability re-run inside those six months is the usual cause of a rejection.[3]
- Filing routine when reliance was available. A WHO-prequalified product filed into the routine queue gives up a published 90-day commitment.[9]
- Sameness drift. A cost-driven excipient or API-source change between the reference filing and the Ethiopian one can take you out of the reliance route.[7]
- Missing the renewal window. Late renewal costs the new-application fee; a missed grace period costs a full new dossier.[4]
Registered. What happens next?
Registration lets the product be placed on the Ethiopian market; it does not move a carton. Every consignment then needs an EFDA import permit, applied for on i-Import by the entity that imports.[2] Who that entity is depends on your buyer: EPSS for public tenders, or a licensed private importer.
Read: how to import medicines from India to Ethiopia, which covers the EPSS tender route, the private-importer route, the import permit, the special import permit and port clearance.
EFDA drug registration: frequently asked questions
EFDA publishes control points rather than a total: acceptance or deficiency notice within 15 days, six months for the applicant to answer queries, and GMP clearance before the certificate. The WHO collaborative procedure carries a 90-day commitment once documents are complete. No end-to-end figure is published for a full assessment of an India-only generic.
Yes. The agent must be physically located in Ethiopia, its technical person should be a pharmacist registered with EFDA, and only that person receives the eRIS login. The agent cannot import the product unless it also holds an EFDA certificate of competence and a trading licence.
No. EFDA’s 2024 reliance guideline lists 39 reference entries, including the US FDA, EMA, MHRA, Health Canada, Swissmedic, TGA, PMDA and WHO prequalification. CDSCO is not among them, so a product approved only in India goes through the full assessment.
Yes, before the marketing authorisation is issued. Under Directive No. 1055/2025 EFDA can inspect on site, run a remote or desk review after a risk assessment, or waive the on-site visit for companies approved by a reference authority. The site must have produced continuously for at least a year.
Five years. Renewal is filed within the 180 days before expiry. A late filing gets a further 180-day grace period at the new-application fee; after that the authorisation is treated as revoked and a full new dossier is needed. Conditional approvals last at most one year.
EFDA’s guidelines point to the Council of Ministers Rate of Service Fees Regulation No. 370/2015 rather than stating amounts. Our registration fee comparator tabulates the dossier evaluation, agent registration and GMP inspection fees from that schedule. Confirm current amounts with EFDA through your agent before budgeting.
eRIS is EFDA’s Electronic Regulatory Information System. i-Register is its product-registration service, where applications, review and marketing authorisation run. i-Import handles import permits and i-License handles certificates of competency for importers, wholesalers and manufacturers.
Only in defined cases. EFDA’s special import permit covers emergencies, verified unmet need or shortage, donations and research; it is reviewed within three working days and valid for one year. It is applied for by the importing institution, case by case, and is not a substitute for registration.[14]
Related on Puizer India
Filing in Ethiopia from an Indian site?
Puizer Cure (OPC) Pvt. Ltd. manufactures tablets and capsules in its WHO-GMP facility in Sonipat, Haryana. Liquids and topicals are made under loan licence at a partner facility, and their site documents are issued for that facility.
Send your product list and the route you expect. We will tell you which products fit a full EFDA assessment and which manufacturer documents we supply for Module 1 and Module 3.
References
- Federal Democratic Republic of Ethiopia. Food and Medicine Administration Proclamation No. 1112/2019, Article 20. Addis Ababa: Federal Negarit Gazeta; 2019. Cited as the legal basis in EFDA’s renewal and non-routine registration guidelines.
- Ethiopian Food and Drug Authority. e-Service: i-Register, i-Import, i-License and eRIS. Addis Ababa: EFDA. Available from: efda.gov.et/e-service. Accessed October 2026.
- Ethiopian Food and Drug Authority. Guideline for Registration of Medicine. EFDA/GDL/017, Version 005. Addis Ababa: EFDA; 30 October 2023. Sections 1.4, 1.6, 1.7, 1.9, 1.10 and General Guidance. Available from: efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Guideline for Renewal of Marketing Authorization. EFDA/GDL/018, Version 003. Addis Ababa: EFDA; 5 December 2023. Sections 2, 3 and 5. Available from: efda.gov.et. Accessed October 2026.
- World Health Organization. Ethiopia achieves major milestone in medicines regulation reaching WHO Maturity Level 3 [news release]. Geneva: WHO; 30 September 2025. Available from: who.int. Accessed October 2026.
- AIM. Mozambique achieves WHO ML3 for having well-functioning regulatory system [trade press]. Maputo; 25 August 2026. Available from: aimnews.org. Accessed October 2026.
- Ethiopian Food and Drug Authority. Guidelines on Reliance for Medicine Marketing Authorization. EFDA/GDL/042, Version 003. Addis Ababa: EFDA; 2 December 2024. Sections 6, 7 and 9; Annex I. Available from: efda.gov.et. Accessed October 2026.
- Council of Ministers. Rate of Service Fees Regulation No. 370/2015. Addis Ababa: EFDA. Available from: efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Guideline for Registration of WHO Pre-qualified Medicines through Collaborative Registration Procedure. EFDA/GDL/021, Version 002. Addis Ababa: EFDA; 30 November 2023. Available from: eris.efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Guideline for Non-Routine Registration. EFDA/GDL/024, Version 001. Addis Ababa: EFDA; 30 November 2023. Citing Directive No. 963/2023, Articles 23 and 24. Available from: efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Ethiopian Essential Medicines List, October 2024 revision. Addis Ababa: EFDA. Available from: efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Medicine Packaging and Labeling Directive No. 1091/2025. Addis Ababa: EFDA; June 2025. Available from: efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Medicine Good Manufacturing Practice Inspection Procedure Directive No. 1055/2025. Addis Ababa: EFDA; April 2025. Articles 4, 5, 9, 10, 11, 13, 18 and 21. Available from: efda.gov.et. Accessed October 2026.
- Ethiopian Food and Drug Authority. Guideline on Medical Products Special Import Permit. EFDA/GDL/051, Version 002. Addis Ababa: EFDA; 2023 (version 002 effective 1 May 2023). Sections 5 to 7. Available from: eris.efda.gov.et. Accessed October 2026.
Disclaimer. Technical and educational content for pharmaceutical professionals; not medical, legal or investment advice. EFDA directives, guidelines and service fees change, and guideline versions are revised without notice. Verify each requirement against the current EFDA publication and with your local agent before acting. Sources were checked on 22 October 2026.